Regulatory Affairs
Regulatory Strategy
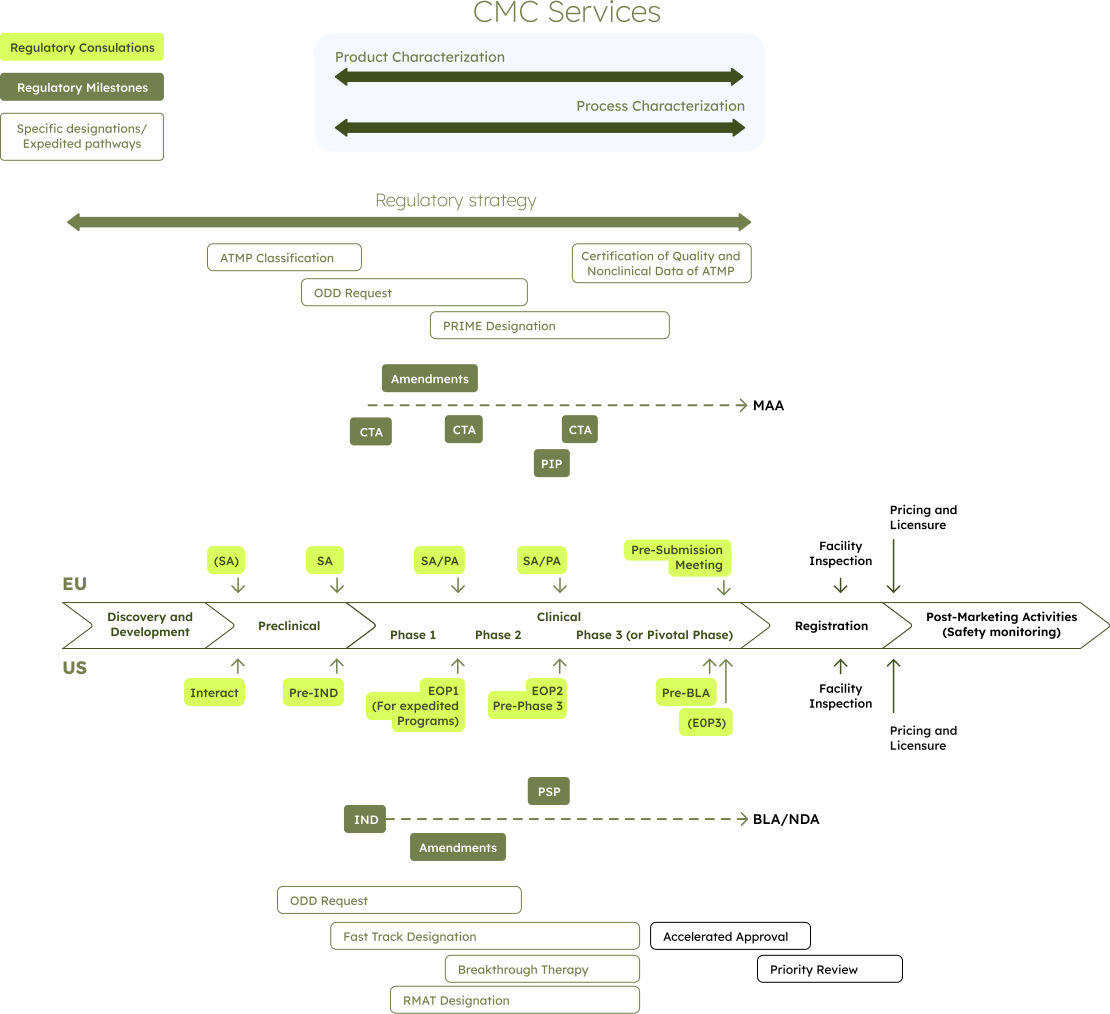
We help you design a global regulatory strategy from the ground up or refine an existing one through a gap assessment, ensuring a clear, actionable plan for CMC and/or nonclinical development – to be validated with regulatory authorities. Our strategic guidance covers the following areas:
- Regulatory Framework
The regulatory landscape is constantly changing. We support you in defining the specific regulatory environment of your product through the determination of product regulatory classification and applicable guidelines based on product type, route of administration, clinical indication, clinical stage and geographical scope. For borderline products, we assist in engaging with regulatory authorities. Securing the right classification is an essential cornerstone for regulatory success. We can also help track your product’s regulatory history, make sure that all recommendations received through scientific advice meetings or the CTA/ IND process are mapped, addressed and incorporated into your strategy. Additionally, we identify potential regulatory risks and develop tailored mitigation strategies. - CMC Strategy
Depending on your product’s development stage, we assist in formulating an initial CMC strategy to establish a solid foundation or conduct a gap analysis of your existing CMC data package or development plan.
See our CMC Service for more details - Preclinical Development plan
Whether you need to define a comprehensive preclinical strategy or assess existing pharmacodynamics, pharmacokinetics, and safety studies, we offer gap analysis and strategic recommendations to optimize your preclinical development.
Regulatory Consultations
We provide expert support in preparing and managing scientific advice (SA)/ protocol assistance (PA) request or (in)formal meetings (INTERACT, pre-IND, EOP (end of phase), and other meeting types) with regulatory authorities (EU-national competent authorities, EMA or US FDA). Early and continuous engagement with competent authorities is key to minimizing regulatory hurdles during drug development.
We assist with:
- Identifying key questions to raise (CMC, Nonclinical, Clinical, Significant benefit, or other strategic considerations)
- Selecting the most appropriate type of advice, agency and optimal timeline
- Authoring or reviewing the Briefing Document/Book to ensure a clear and compelling regulatory case
- Managing submission and communications with Authorities throughout the process
- Preparing and/or participating to meeting with the Agency, ensuring your team is well-equipped
- Drafting or reviewing meeting minutes for regulatory compliance
- Interpreting the Final Advice and implementing recommendations into your drug development plan/ regulatory strategy
Note: For formal FDA meetings, we need to partner with a US-based company
Regulatory Milestones
We provide hands-on support throughout key regulatory milestones in your drug clinical development:
- IMPD (Investigational Medicinal Product Dossier) (EU) or CTD Module 3 (US):
- Full drafting, critical review, and amendment preparation (including compilation of the changes with rationale)
- Assistance with responding to CMC-related regulatory queries
- IB (Investigator’s Brochure):
- Development from scratch or expert review
- Updates and amendments with detailed justification for changes
- Support in responding to nonclinical and clinical regulatory questions
- CTA (Clinical Trial Application) (EU):
- Preparation, submission, and full management through the CTIS (Clinical Trials Information System)
Note: For IND application (US), we need to partner with a US-based company.
- PIP (Paediatric Investigation Plan) (EU) / iPSP (initial Pediatric Study Plan) (US):
- Drafting, submission, responses to request for modifications, and maintenance of the agreed initial plan
Note: For the iPSP, we need to partner with a US-based company.
Specific Designations/ Expedited Pathways
We provide support for the preparation and submission of the following requests to the European Medicines Agency (EMA) or to the U.S. Food and Drug Administration (FDA):
- Advanced Therapy Medicinal Product (ATMP) classification (EMA)
- Orphan Drug Designation (ODD) (EMA & FDA)
- PRIME (Priority Medicines) designation (EMA)
- Certification of quality and/or nonclinical data for ATMPs (EMA, for SMEs only)
- Fast Track, RMAT (Regenerative Medicine Advanced Therapy), Breakthrough Therapy designations (FDA)
Note: For US requests, we need to partner with a US-based company
Looking for Specialized Regulatory CMC Expertise?
See our CMC Services for more detailsschema-01